Explorar las categorías
Español
$
USD
Conéctate con tu IA
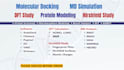
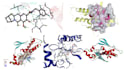
Expert Molecular Docking, MD Simulation & Protein Modeling
Looking for accurate computational support for your research or drug discovery project? Youre in the right place.
I am Rahul Singh, a Research Scholar with 3+ years of experience in bioinformatics and computational chemistry, specializing in in-silico drug design and molecular modeling.
Services:
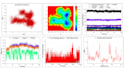
Analysis Includes:
RMSD, RMSF, SASA, Radius of Gyration, Hydrogen Bonds, PCA, MM/PBSA, GBSA, Protein-Ligand Interactions
Why Choose Me?
Please message before placing an order to discuss your project.
ORCID: https://orcid.org/0009-0000-0994-8912
Thank You
Rahul Singh
Informa al freelancer cualquier preferencia o inquietud que tengas con respecto al uso de herramientas de IA en la finalización y/o entrega de tu pedido.
Professional Bioinformatician Expert in Drug Design and Data Analysis
Idiomas


